GLP products and FDA compliance

August 22, 2025

Understanding GLPs in Relation to FDA Standards

Good Laboratory Practices (GLP) are critical for ensuring the integrity and quality of nonclinical study data submitted to the FDA. Governing factors, such as GLP regulations, play an instrumental role in maintaining the reliability of safety assessments necessary for product approvals across a variety of FDA-regulated industries. This narrative explores the multiple dimensions of GLP and FDA compliance, providing insights into their interconnected framework.

GLP Regulations and Their Relationship with FDA Compliance

Overview of GLP regulations under the FDA

Good Laboratory Practice (GLP) regulations, as outlined in 21 CFR Part 58, are critical for nonclinical laboratory studies. These regulations apply to laboratory studies supporting applications for research or marketing permits for products regulated by the FDA, including drugs, medical devices, and food additives.

Regulatory compliance under GLP is not only about following procedures; it encompasses the entire framework within which research is conducted, assuring that studies are organized, personnel are qualified, and facilities are adequate. This rigorous approach helps ensure that the data produced is reliable and meets scientific standards required for FDA review.

Importance of GLP compliance for data integrity

Compliance with GLP is vital for maintaining the quality and integrity of safety data. The FDA mandates adherence at every phase of nonclinical studies, and the role of Quality Assurance Units (QAU) is crucial. These units are tasked with monitoring compliance and validating that study reports accurately represent the work performed, thereby safeguarding data integrity crucial for FDA applications.

Being compliant with GLP helps mitigate risks of noncompliance, which can bring about civil monetary penalties or increased scrutiny during FDA inspections.

Accountability and documentation required under GLP

GLP regulations necessitate extensive documentation that serves to ensure accountability and traceability of studies. This includes detailed study protocols, reporting of results, and comprehensive record-keeping for at least two years post-FDA approval of the relevant application. Such meticulous documentation is not only vital for regulatory compliance but also supports the reconstruction and validation of study findings, reinforcing trust in the data presented to the FDA.

The Role of Quality Assurance in GLP Compliance

What is the role of quality assurance in GLP compliance?

Quality assurance (QA) is crucial for maintaining Good Laboratory Practice (GLP) compliance by ensuring the integrity of laboratory practices and the accuracy of study data. In GLP laboratories, QA units provide independent oversight, conducting thorough audits and inspections to confirm adherence to established standards.

These independent teams are responsible for implementing systematic quality checks throughout all phases of nonclinical laboratory studies. This includes developing and maintaining detailed documentation, such as Standard Operating Procedures (SOPs), which guide laboratory processes to minimize variability.

Moreover, QA professionals play a significant role in training laboratory personnel on GLP standards. They facilitate workshops and training sessions that reinforce the importance of compliance, thus fostering an environment where reliability and quality are prioritized.

QA units not only enhance procedural integrity but also provide management with confidence in the validity of the data submitted to regulatory authorities. Together, these efforts ensure the laboratory's operations align with regulatory expectations and help safeguard public health by yielding dependable safety data for products such as drugs and medical devices.

Understanding the FDA's Inspection Processes for GLP Studies

What is the FDA's inspection process related to GLP studies?

The FDA's inspection process for Good Laboratory Practice (GLP) studies is comprehensive and aims to ensure compliance with regulations outlined in Part 58 of Title 21 of the Code of Federal Regulations.

Frequency and nature of FDA inspections

Inspections typically occur every two years. These inspections involve a thorough assessment of laboratory facilities, personnel training, and organizational structures. Inspectors look closely at study performance, reviewing essential documents such as the Master Schedule Sheet and final reports.

Evaluation criteria during inspections

During the inspection, the FDA evaluates various aspects including:

  • Data integrity
  • Adherence to documented study protocols
  • Oversight of the Quality Assurance Unit (QAU) that monitors compliance

FDA-483 forms and post-inspection procedures

If any objectionable practices are identified, they are documented via FDA-483 forms. After an inspection, laboratories are required to address significant findings promptly. They must demonstrate commitment to implementing corrective and preventive measures to maintain compliance with GLP standards, ensuring the reliability of safety data submitted to the FDA.

Key Responsibilities and Legal Implications of GLP Noncompliance

What responsibilities and legal ramifications are associated with noncompliance of GLPs?

Noncompliance with Good Laboratory Practices (GLPs) attracts significant responsibilities and legal ramifications. Laboratories are required to report any GLP deviations within their FDA submissions. This lack of adherence can lead to the rejection of crucial safety data, ultimately jeopardizing regulatory approvals.

Additionally, failure to conform to GLP standards may expose facilities to civil or even criminal penalties. Such actions are enforceable in U.S. District Courts and could infringe on laws such as the Toxic Substances Control Act (TSCA). This presents a risk for product approvals and extends liability to the testing facility.

What are the consequences of refusing an inspection by the FDA?

Refusal to permit an FDA inspection can have severe consequences. When laboratories deny access for inspections, any associated study data may be rendered unreliable. This impacts the integrity of the laboratory’s submissions and affects the credibility of ongoing or future research projects.

How should labs manage GLP deviations?

It’s crucial for labs to have a management strategy in place for GLP deviations. Laboratories should document and report these deviations promptly, as outlined by FDA regulations. This approach ensures accountability and helps maintain compliance, which is vital for upholding the reliability of safety data necessary for regulatory decision-making.

Economic Impact of GLP Regulations on Laboratory Operations

How do GLP regulations impact the economics of laboratory operations?

GLP regulations significantly influence laboratory operation economics by imposing added costs for compliance, quality assurance, and the creation of Standard Operating Procedures (SOPs). The estimated annual costs for implementing a GLP Quality System are projected to be around $51.9 million , adjusting for a 3% discount rate over a decade. These costs encompass extensive reporting and recordkeeping obligations that facilities must adhere to.

In addition to these ongoing expenses, there are one-time costs for initial compliance assessments and necessary staff training to align with GLP standards. Inspections and compliance monitoring by regulatory bodies also contribute to operational expenses.

However, on the positive side, GLP compliance can lead to improved operational efficiency. By ensuring the integrity and quality of data, laboratories can enhance scientific reliability, ultimately leading to more effective and innovative research outcomes. This rigor can boost return on investment and facilitate faster product development cycles in the pharmaceutical industry.

Moreover, GLP regulations impact wider economic aspects, particularly in the pharmaceutical sector . For instance, the growth of GLP-1 medications is expected to reduce healthcare spending significantly, affecting over 50 industries by shifting spending patterns.

Thus, while GLP compliance increases operational costs, it also promotes efficiency and can foster economic growth in related markets, presenting a dual-edged impact on laboratory economics.

Scope and Specific Provisions of GLP Regulations for Nonclinical Studies

Requirements under 21 CFR Part 58

Good Laboratory Practice (GLP) regulations, as outlined in 21 CFR Part 58, are pivotal for nonclinical laboratory studies that support applications for products regulated by the FDA, including drugs and medical devices. Compliance requires that these studies are planned, performed, and reported with integrity and adherence to established protocols. Applicants must provide a compliance statement confirming their adherence to GLP provisions.

Study Integrity and Data Quality Assurance

To ensure the integrity of GLP studies, the FDA mandates thorough inspections during all phases of the research. These inspections verify that studies are adequately designed and executed, producing reliable data for FDA review. Moreover, quality assurance measures are essential; facilities must have a dedicated quality assurance unit to monitor compliance, report findings, and ensure adherence to standard operating procedures (SOPs).

Detailed Scope, Including Test Article Regulations

GLP regulations cover a wide range of test articles, defined as any product subject to FDA regulation, encompassing medical devices, drugs, and food additives. Nonclinical studies must accurately characterize test and control articles while maintaining accountability in their methodologies. Facilities conducting such studies are required to retain records of compliance for a minimum of two years post-approval to uphold transparency and data integrity.

Current Issues Impacting GLP-1 Medication Availability

Status of GLP-1 Medication Shortages

As of October 02, 2024, the shortage of tirzepatide injection, a prominent GLP-1 medication, has been resolved. This follows a period of increased demand beginning in 2022, which impacted availability for patients needing this treatment.

Impact of Resolved and Ongoing Shortages

While tirzepatide is now back in stock, issues with other GLP-1 medications, specifically dulaglutide and semaglutide injections, persist. These ongoing shortages highlight the complexities involved in maintaining supply for high-demand treatments in the market.

FDA's Regulatory Stance on GLP-1 Medications

The FDA remains vigilant about the market status of GLP-1 medications. They have reiterated that compounded versions of these drugs, unlike FDA-approved treatments, do not undergo the same rigorous safety and efficacy evaluations. This raises concerns about the potential risks patients may face, particularly with doses that differ from established standards.

To ensure safe usage, it is crucial for patients to engage with healthcare professionals regarding their treatment options, focusing on FDA-approved remedies such as:

Medication Name Brand Name Administration Method Use Case
Dulaglutide Trulicity® Subcutaneous injection Type 2 diabetes and obesity
Exenatide Byetta® Subcutaneous injection Type 2 diabetes
Exenatide extended-release Bydureon® Subcutaneous injection Type 2 diabetes
Liraglutide Victoza® Subcutaneous injection Type 2 diabetes and obesity
Lixisenatide Adlyxin® Subcutaneous injection Type 2 diabetes
Semaglutide injection Ozempic® Subcutaneous injection Type 2 diabetes and obesity
Semaglutide tablets Rybelsus® Oral tablet Type 2 diabetes

Each of these GLP-1 medications offers distinct benefits for managing conditions like Type 2 diabetes and obesity, emphasizing the necessity for FDA oversight to ensure patient safety.

Regulatory Framework and the Role of GLP in FDA Compliance Programs

FDA's Role in GLP Enforcement

The implementation of Good Laboratory Practice (GLP) regulations is primarily enforced by the FDA to ensure that nonclinical laboratory studies meet the necessary standards for data quality and integrity. This enforcement is crucial for products regulated by the FDA, including drugs and medical devices. Regular inspections of laboratories are conducted to confirm adherence to GLP standards, and noncompliance can result in significant penalties, including the exclusion of studies from FDA application support.

Title 21 CFR Part 58 Implications

Title 21 CFR Part 58 establishes specific regulatory criteria that must be followed during nonclinical studies. It underscores the importance of compliance with GLP, requiring comprehensive documentation and the implementation of a Quality Assurance Unit (QAU). These regulations aim to maintain rigorous standards throughout all phases of a study, ensuring reliable safety data is submitted for FDA review.

GLP Framework Accountability

The GLP framework promotes accountability in laboratory practices by detailing responsibilities for study directors, QA protocols, and maintaining accurate records. It ensures that every phase of a nonclinical study is inspected and monitored, thus enhancing the overall reliability of the data generated for regulatory submissions. By adhering to GLPs, laboratories can significantly contribute to the safety and efficacy evaluations conducted by the FDA.

Educational Resources for Effective GLP Compliance

What educational resources are available for GLP compliance training?

There are a variety of educational resources dedicated to GLP compliance training designed for laboratories and individuals involved in nonclinical research. These resources include:

  • Online Courses : Comprehensive courses such as the Basic Training: Good Laboratory Practice cover essential GLP regulations mandated by the FDA, EPA, and OECD.
  • Webinars : These are often interactive sessions that offer insights into specific GLP topics, featuring expert speakers from the industry.
  • Industry Workshops : Hands-on workshops provide practical training, allowing participants to apply GLP principles in a controlled environment.

Importance of training for maintaining GLP standards

Training is paramount in upholding GLP standards. It equips participants with the necessary knowledge to ensure compliance during nonclinical studies. Key benefits of effective training include:

  • Understanding GLP guidelines thoroughly, including history and regulations.
  • Preparing for regulatory inspections, minimizing the risk of non-compliance.
  • Ensuring that quality assurance protocols are followed, resulting in reliable data.

Targeted audiences and certification processes

The targeted audiences for GLP training include:

  • Scientists : Engaged in conducting nonclinical studies.
  • Quality Assurance Personnel : Responsible for monitoring compliance.
  • Study Directors : Who oversee laboratory practices.

Moreover, many training programs offer certification processes, allowing participants to obtain Continuing Education (CE) credits upon completion. This not only recognizes their proficiency in GLP standards but also enhances their professional credibility in the industry.

Through these educational resources, organizations can enhance their compliance with Good Laboratory Practices, thereby contributing to the integrity of safety studies supporting FDA-regulated products.

The Vital Role of GLP in Product Safety and FDA Compliance

In the realm of nonclinical laboratory studies, compliance with Good Laboratory Practices is indispensable for maintaining the integrity, reliability, and acceptability of safety data in support of FDA applications. With rigorous inspection processes, substantial compliance responsibilities, and high economic stakes, GLP adherence is a complex yet critical obligation for laboratories. Educational resources and strategic implementation of quality systems further strengthen the framework, ultimately safeguarding the efficacy of regulated products and promoting public health. Understanding and navigating this landscape is key for laboratories aiming to achieve excellence in regulatory submissions and contribute to the continuous advancement of safety standards in product development.

References

Person living with end stage renal disease going about an ordinary morning at home
September 1, 2026
Straight answers to what people on dialysis ask most: nutrition, phosphate, energy, travel, and the emotional weight of ongoing treatment.
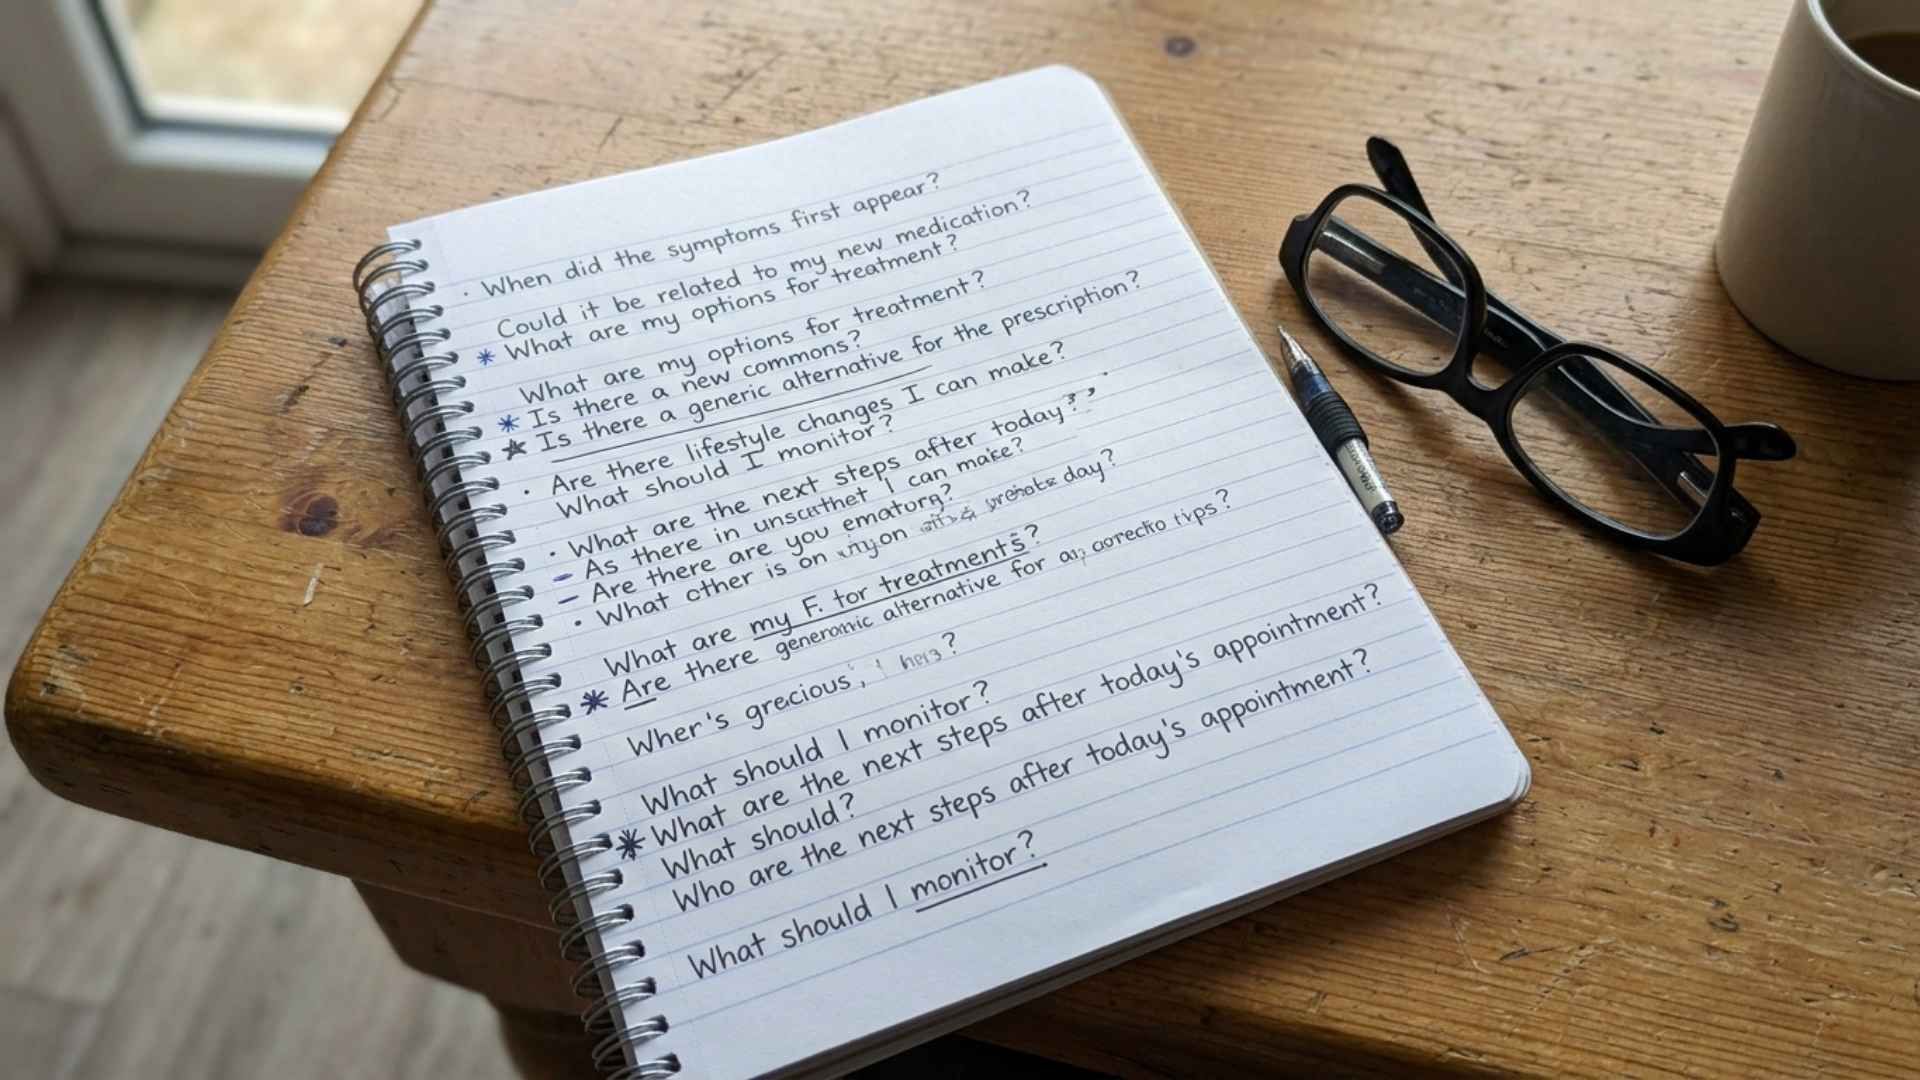
Handwritten list of questions prepared for a dialysis appointment about IDPN therapy
September 1, 2026
The questions worth bringing to your nephrologist or dietitian before starting intradialytic parenteral nutrition, grouped and ready to print.
Adult daughter and her mother reviewing a delivery schedule together in the kitchen at home
September 1, 2026
A practical guide for caregivers: what your role is, what the pharmacy can share with you, what to keep track of, and how to protect your own energy.
Family sharing a meal at home while one member on parenteral nutrition joins the table
September 1, 2026
Whether you can eat on parenteral nutrition, why appetite and taste change, and how to handle meals with family while receiving IV nutrition.
Suitcase packed with a small cooler and paperwork for traveling while on parenteral nutrition
August 31, 2026
How to plan a trip on TPN or home parenteral nutrition: supplies, refrigeration, air travel documentation, and how far ahead to call your pharmacy.
Home delivery of TPN supplies unpacked on a kitchen table during the first week of therapy
August 31, 2026
A day-by-day look at starting total parenteral nutrition at home: the first delivery, the first infusion, night one, and when to call for help.
Dialysis center staff coordinating phosphate binder supply with a pharmacy partner
August 11, 2026
What dialysis centers should look for in a phosphate binder supply partner: reliable delivery, formulary breadth, insurance support, and coordination.
Renal dietitian reviewing phosphate levels with a dialysis patient
August 11, 2026
How hyperphosphatemia is managed in ESRD dialysis patients, diet, dialysis, and phosphate binders, and why reliable binder supply keeps therapy on track.
July 29, 2026
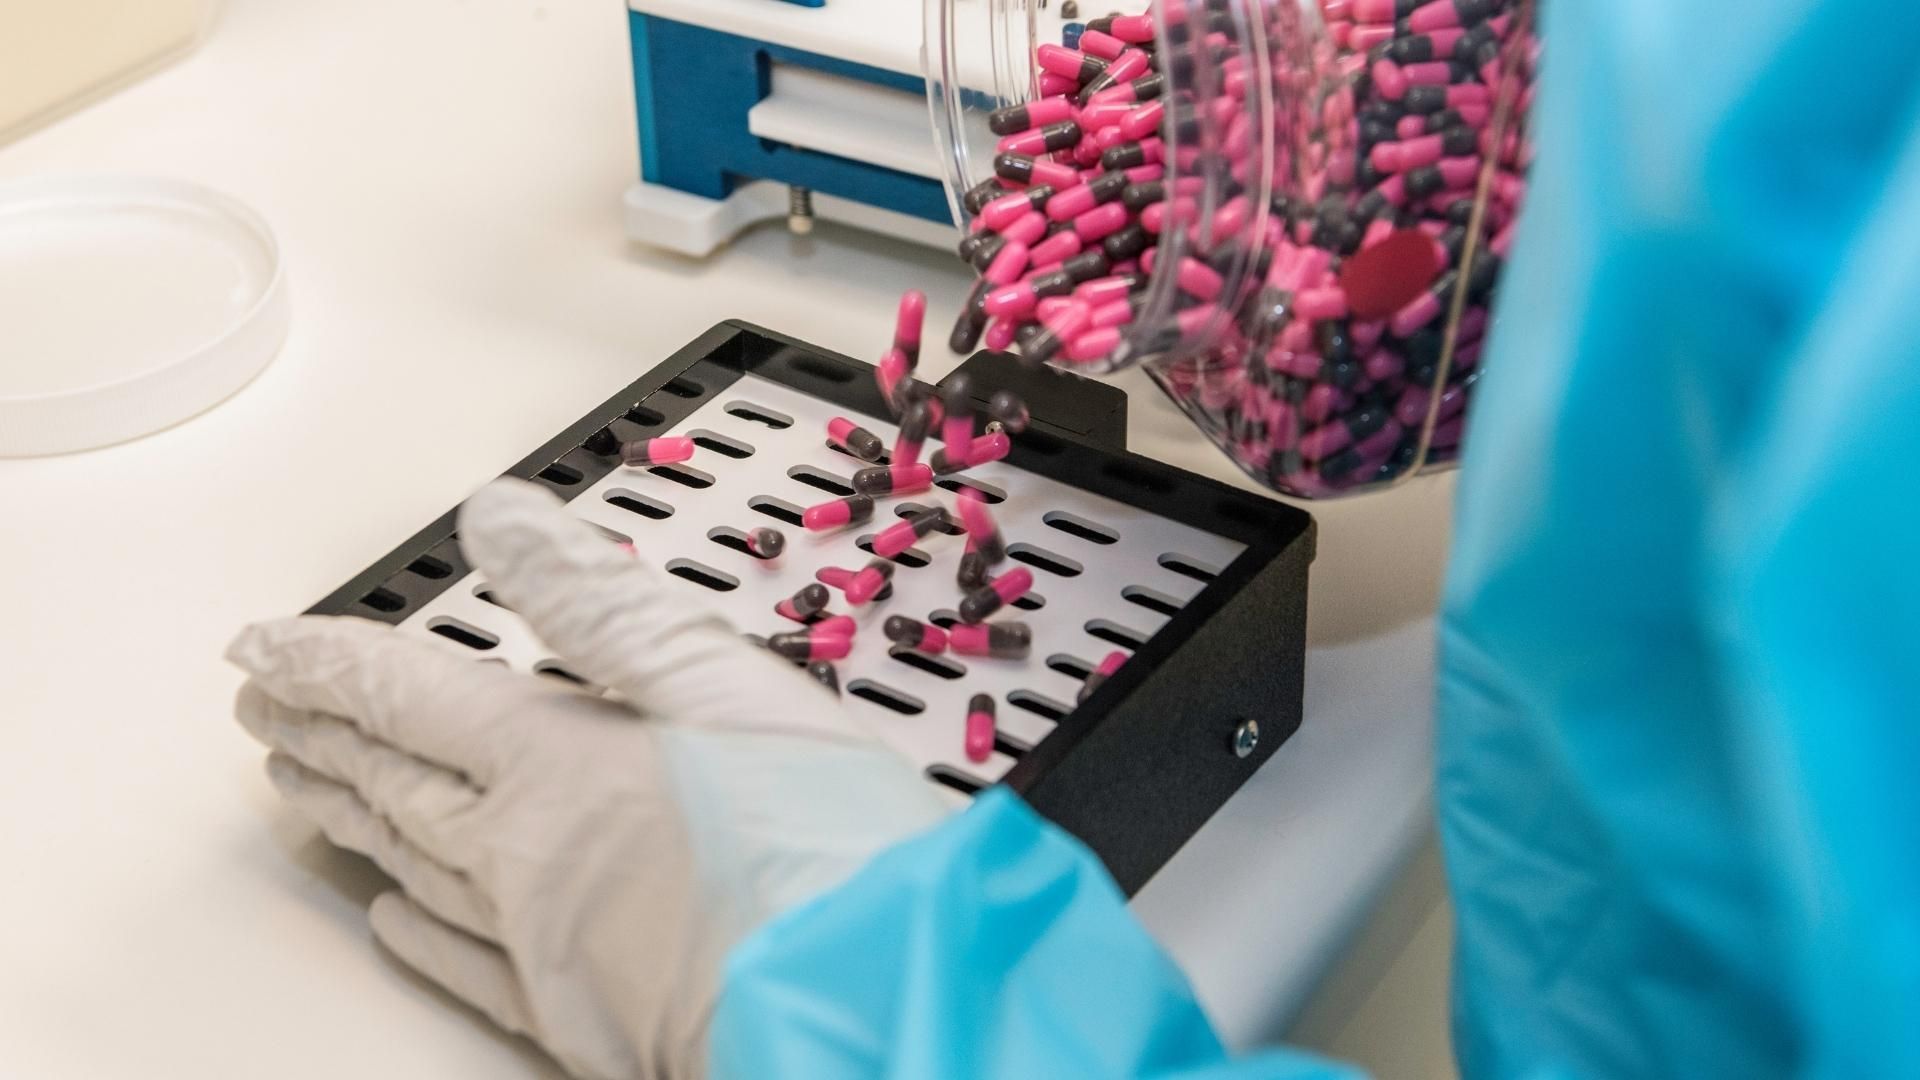
Compounded medications are a routine part of modern care, but unlike mass-manufactured drugs, each one is prepared individually, which means the standards behind its preparation carry real weight. For providers who order or refer for compounded therapies, a working understanding of those standards is the clearest way to judge whether a pharmacy is safe to trust. This overview covers the standards that govern sterile compounding and what they mean for the medications your patients receive. Why Standards Matter More in Compounding A commercially manufactured drug is produced under FDA oversight in large, validated batches. A compounded medication is prepared for an individual patient, often in small quantities, sometimes because no commercial equivalent exists. That flexibility is exactly why it's clinically valuable — and why the standards around preparation exist. Without rigorous controls, the same customization that helps patients could introduce contamination or error. Our guide to our sterile compounding pharmacy describes how those controls come together in practice. The Regulatory Landscape Sterile compounding sits within a layered framework: The United States Pharmacopeia (USP) publishes the general chapters, most importantly <797> and <800>, that define how compounded preparations should be made and handled. State boards of pharmacy license and inspect pharmacies, including where they're permitted to ship. Multi-state operations must maintain licensure in each state they serve. Accreditation bodies (ACHC, NABP, PCAB) provide independent verification that a pharmacy meets recognized standards. Together, these create the accountability structure behind every compounded dose. USP <797>: Sterile Preparations USP <797> is the cornerstone standard for compounded sterile preparations. It governs the conditions and practices required to keep a preparation free of contamination, including personnel training and garbing, ISO-classified air quality and engineering controls, environmental monitoring, and beyond-use dating. For any medication infused directly into the body, <797> compliance is the baseline expectation, not a premium feature. USP <800>: Hazardous Drugs USP <800> adds a containment layer for hazardous drugs, protecting personnel, patients, and the environment from unintended exposure. It specifies containment engineering controls, PPE, and validated handling procedures for drugs identified as hazardous. When a preparation is both sterile and hazardous, it must satisfy <797> and <800> together. For the details, see our breakdown of USP <800> requirements . Accreditation: Independent Verification Standards are only meaningful if someone checks that they're being met. Accreditation from ACHC, NABP, and PCAB means a pharmacy has been inspected against national benchmarks by an outside authority. For a provider, accreditation is a shortcut to confidence: it signals that the pharmacy's claims about its processes have been independently validated. The Quality-Control Process Behind the standards is a repeatable process that governs each preparation: Pharmacist review of the prescription for appropriateness, compatibility, and dosing Preparation by trained technicians under aseptic conditions In-process and final verification by a licensed pharmacist Sterility testing or filtration validation where required This chain is what turns a standard on paper into a safe medication in a patient's hands. What This Means When You Order When you refer a patient for a compounded therapy, you're effectively extending your own standard of care into another facility. Knowing that a pharmacy is USP-compliant, accredited, and pharmacist-led lets you refer with confidence that the medication will meet the same expectations you'd hold for anything you administer yourself. If you're weighing options, our guide on how to choose a compounding partner walks through the criteria. The Pharmko Standard Pharmko compounds every TPN, IDPN, and IPN preparation in a USP <797>-compliant, ISO Class 5 cleanroom, under ACHC, NABP, and PCAB accreditation, with a licensed pharmacist reviewing and releasing each formula. For the patients who receive our preparations, many of them managing End Stage Renal Disease (ESRD), that standard is the point. Refer a Patient → · [1 (877) 540-2003]
Physician evaluating a sterile compounding pharmacy partner
July 29, 2026
How to evaluate a sterile compounding pharmacy: accreditation, USP compliance, clinical support, turnaround, and the questions every practice should ask.
Show More